Featured Publications
All Publications

The growing field of data-driven materials research poses a challenge to educators: teaching machine learning to materials scientists. We share our recent experiences and lessons learnt from organizing educational sessions at the fall 2021 meeting of the Materials Research Society. READ MORE
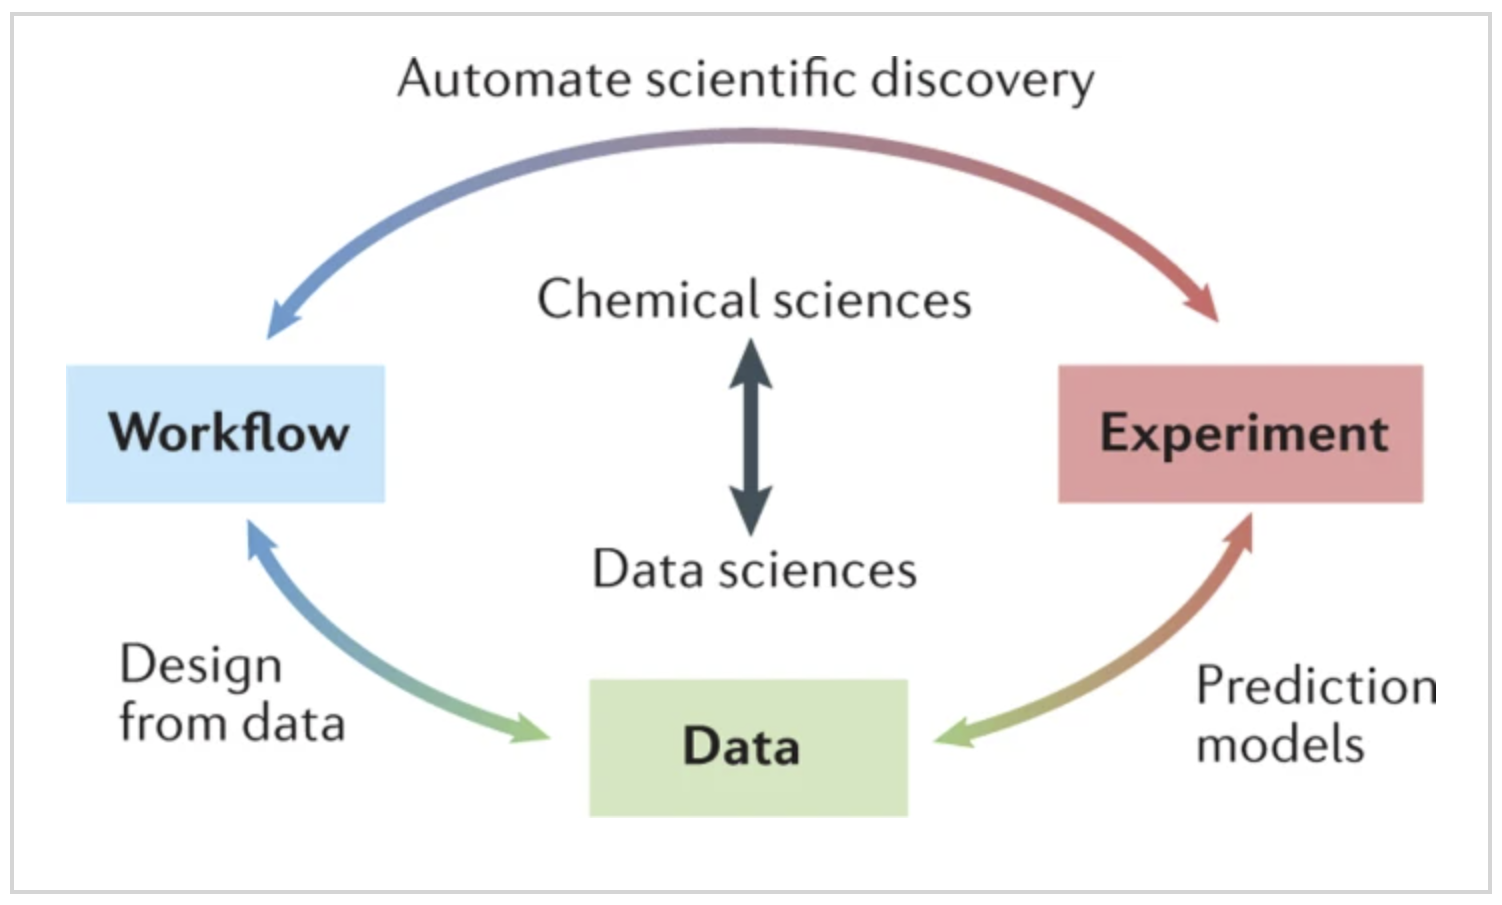
The physical sciences community is increasingly taking advantage of the possibilities offered by modern data science to solve problems in experimental chemistry and potentially to change the way we design, conduct and understand results from experiments. Successfully exploiting these opportunities involves considerable challenges. In this Expert Recommendation, we focus on experimental co-design and its importance to experimental chemistry. We provide examples of how data science is changing the way we conduct experiments, and we outline opportunities for further integration of data science and experimental chemistry to advance these fields. Our recommendations include establishing stronger links between chemists and data scientists; developing chemistry-specific data science methods; integrating algorithms, software and hardware to ‘co-design’ chemistry experiments from inception; and combining diverse and disparate data sources into a data network for chemistry research. READ MORE
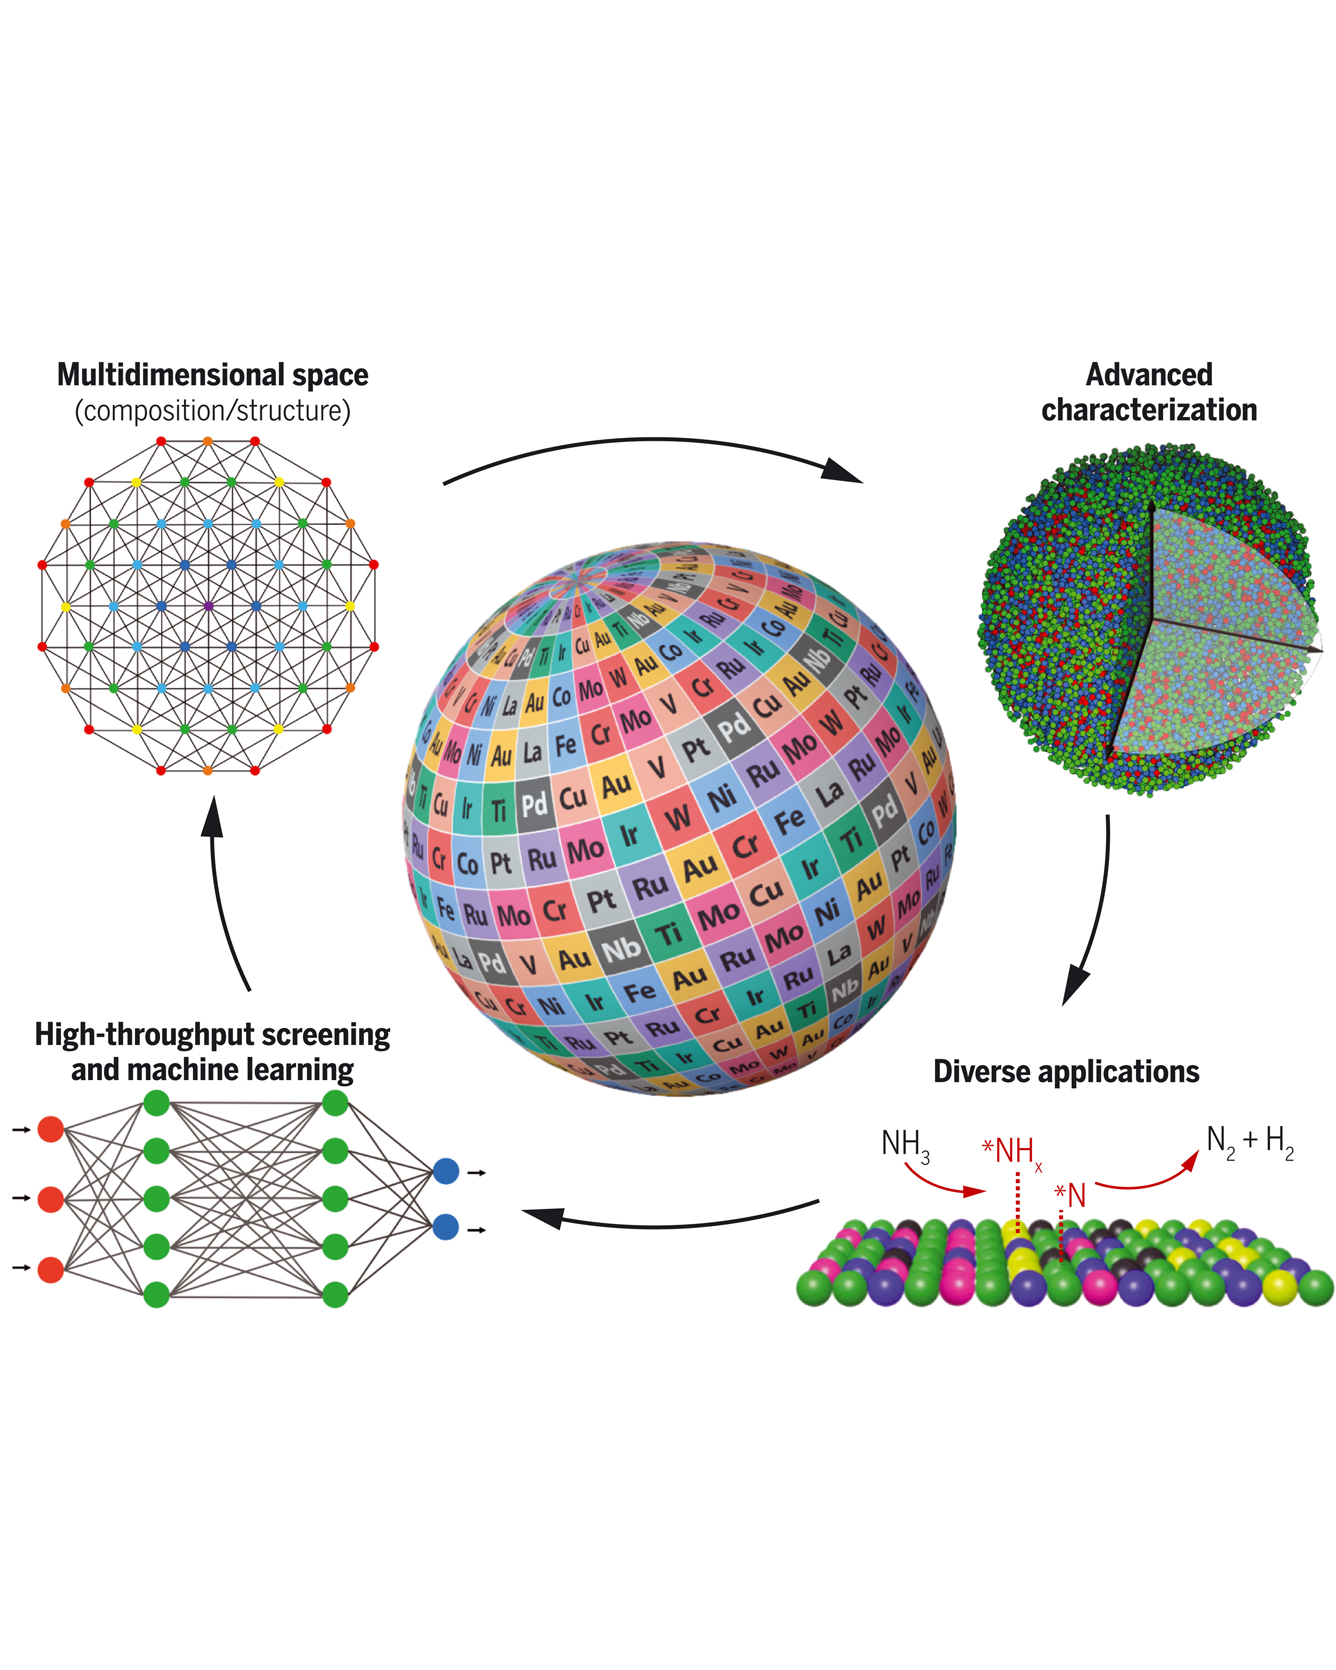
High-entropy nanoparticles have become a rapidly growing area of research in recent years. Because of their multielemental compositions and unique high-entropy mixing states (i.e., solid-solution) that can lead to tunable activity and enhanced stability, these nanoparticles have received notable attention for catalyst design and exploration. However, this strong potential is also accompanied by grand challenges originating from their vast compositional space and complex atomic structure, which hinder comprehensive exploration and fundamental understanding. Through a multidisciplinary view of synthesis, characterization, catalytic applications, high-throughput screening, and data-driven materials discovery, this review is dedicated to discussing the important progress of high-entropy nanoparticles and unveiling the critical needs for their future development for catalysis, energy, and sustainability applications. READ MORE
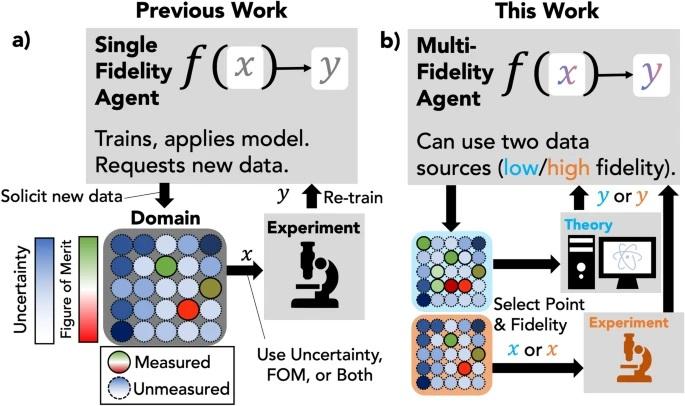
Sequential learning for materials discovery is a paradigm where a computational agent solicits new data to simultaneously update a model in service of exploration (finding the largest number of materials that meet some criteria) or exploitation (finding materials with an ideal figure of merit). In real-world discovery campaigns, new data acquisition may be costly and an optimal strategy may involve using and acquiring data with different levels of fidelity, such as first-principles calculation to supplement an experiment. In this work, we introduce agents which can operate on multiple data fidelities, and benchmark their performance on an emulated discovery campaign to find materials with desired band gap values. The fidelities of data come from the results of DFT calculations as low fidelity and experimental results as high fidelity. We demonstrate performance gains of agents which incorporate multi-fidelity data in two contexts: either using a large body of low fidelity data as a prior knowledge base or acquiring low fidelity data in-tandem with experimental data. This advance provides a tool that enables materials scientists to test various acquisition and model hyperparameters to maximize the discovery rate of their own multi-fidelity sequential learning campaigns for materials discovery. This may also serve as a reference point for those who are interested in practical strategies that can be used when multiple data sources are available for active or sequential learning campaigns. READ MORE
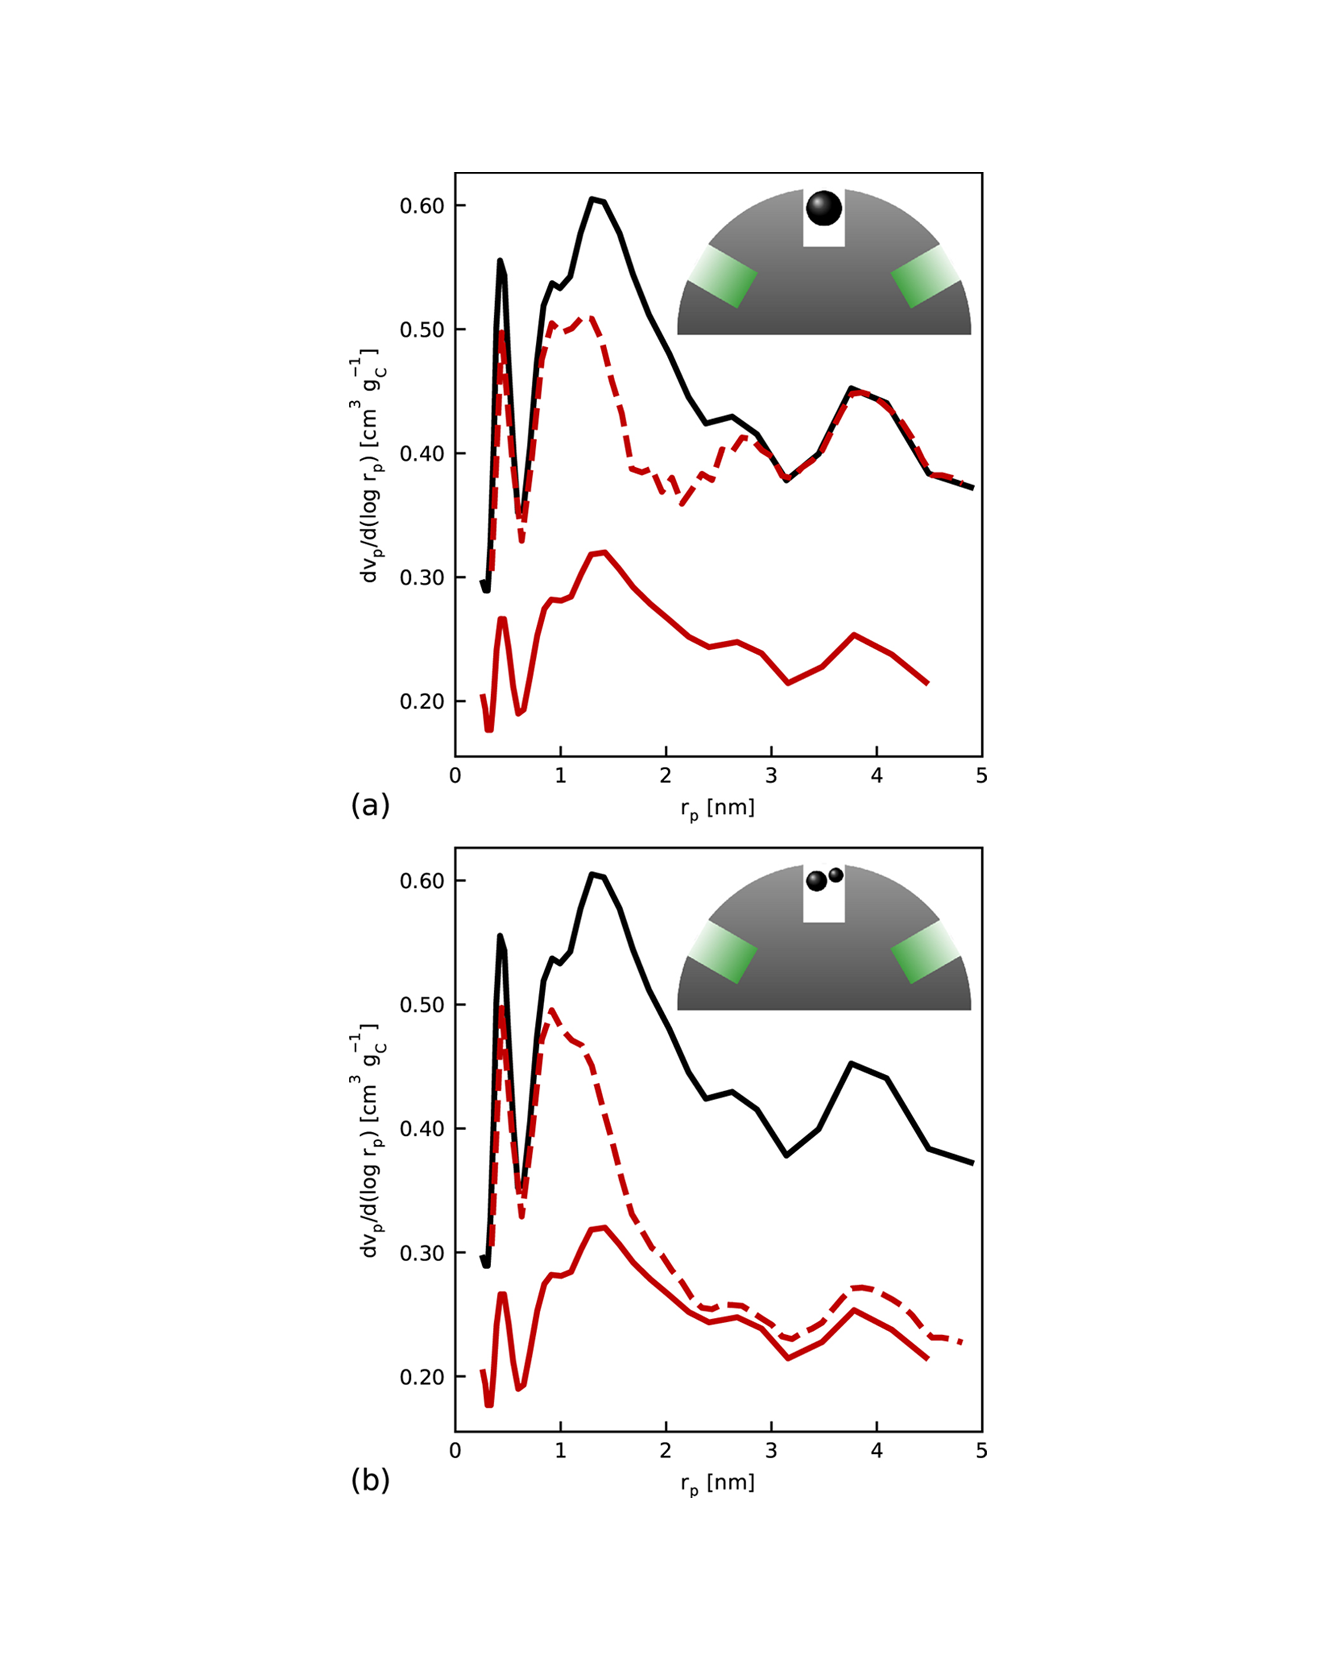
We present a model of the cathode catalyst layer morphology before and after loading a porous catalyst support with Pt and ionomer. Support nanopores and catalyst particles within pores and on the support surface are described by size distributions, allowing for qualitative processes during the addition of a material phase to be dependent on the observed pore and particle size. A particular focus is put on the interplay of pore impregnation and blockage due to ionomer loading and the consequences for the Pt/ionomer interface, ionomer film thickness and protonic binding of particles within pores. We used the model to emulate six catalyst/support combinations from literature with different porosity, surface area and pore size distributions of the support as well as varying particle size distributions and ionomer/carbon ratios. Besides providing qualitatively and quantitatively accurate predictions, the model is able to explain why the protonically active catalyst surface area has been reported to not increase monotonically with ionomer addition for some supports, but rather decrease again when the optimum ionomer content is exceeded. The proposed model constitutes a fast translation from manufacturing parameters to catalyst layer morphology which can be incorporated into existing performance and degradation models in a straightforward way. READ MORE
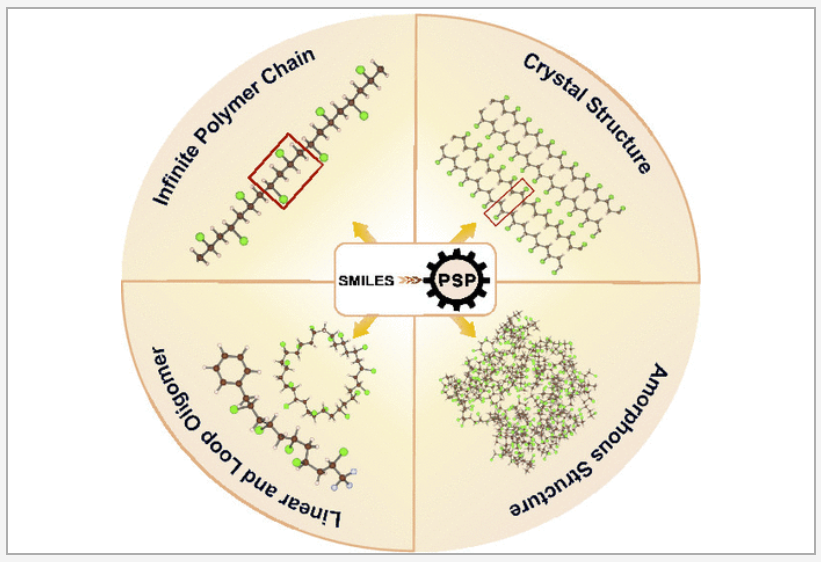
Three-dimensional atomic-level models of polymers are the starting points for physics-based simulation studies. A capability to generate reasonable initial structural models is highly desired for this purpose. We have developed a python toolkit, namely, polymer structure predictor (PSP), to generate a hierarchy of polymer models, ranging from oligomers to infinite chains to crystals to amorphous models, using a simplified molecular-input line-entry system (SMILES) string of the polymer repeat unit as the primary input. This toolkit allows users to tune several parameters to manage the quality and scale of models and computational cost. The output structures and accompanying force field (GAFF2/OPLS-AA) parameter files can be used for downstream ab initio and molecular dynamics simulations. The PSP package includes a Colab notebook where users can go through several examples, building their own models, visualizing them, and downloading them for later use. The PSP toolkit, being a first of its kind, will facilitate automation in polymer property prediction and design. READ MORE
Chapter 3
The materials discovery process naturally presents a slew of questions about the character of a material. These range from simply trying to learn basic facts about the atomic structure of the compound [1–3] all the way to producing a time-resolved profile of a functional process in operando from start to finish [4]. Spectros-copy is the process of measuring a materials’ response to external electromagnetic stimulus to deduce the properties of interest. Spectroscopy can help to solve problems in experimental design and decision-making (probing response of certain energy domains, measuring particular properties), inference (moving from raw data to the property of interest), and analysis (rationalizing and interpreting the data). These are all classes of problems which artificial intelligence (AI) is well-posed to address.As such, the interface between practitioners in both spectroscopy and AI has sparked a great deal of excitement and research activity. Because characterization is a crucial process of the materials discovery pipeline, insightful pairings of algorithms and experimental protocols can lead to gains in both experimental efficiency and accuracy.These gains may significantly compress the timescales of experimental procedures and help experimentalists learn more in less time or answer questions which were previously inaccessible with a given experimental apparatus. READ MORE
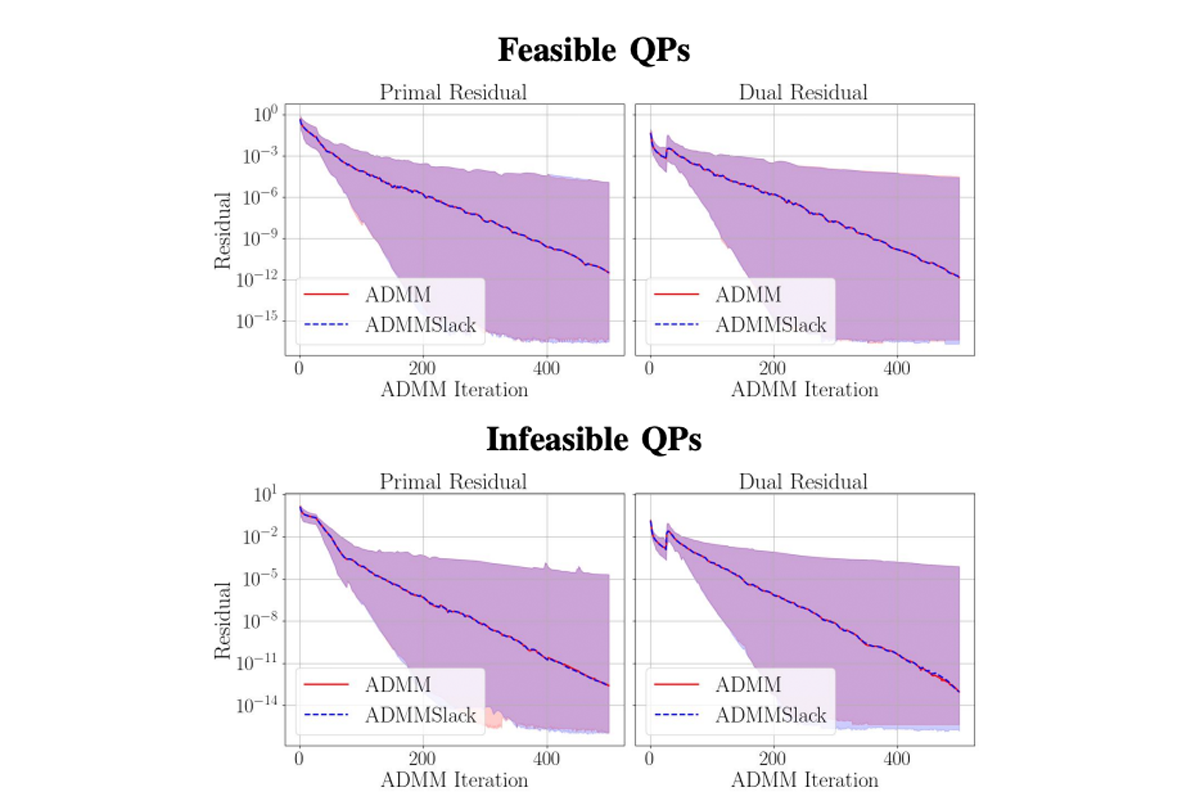
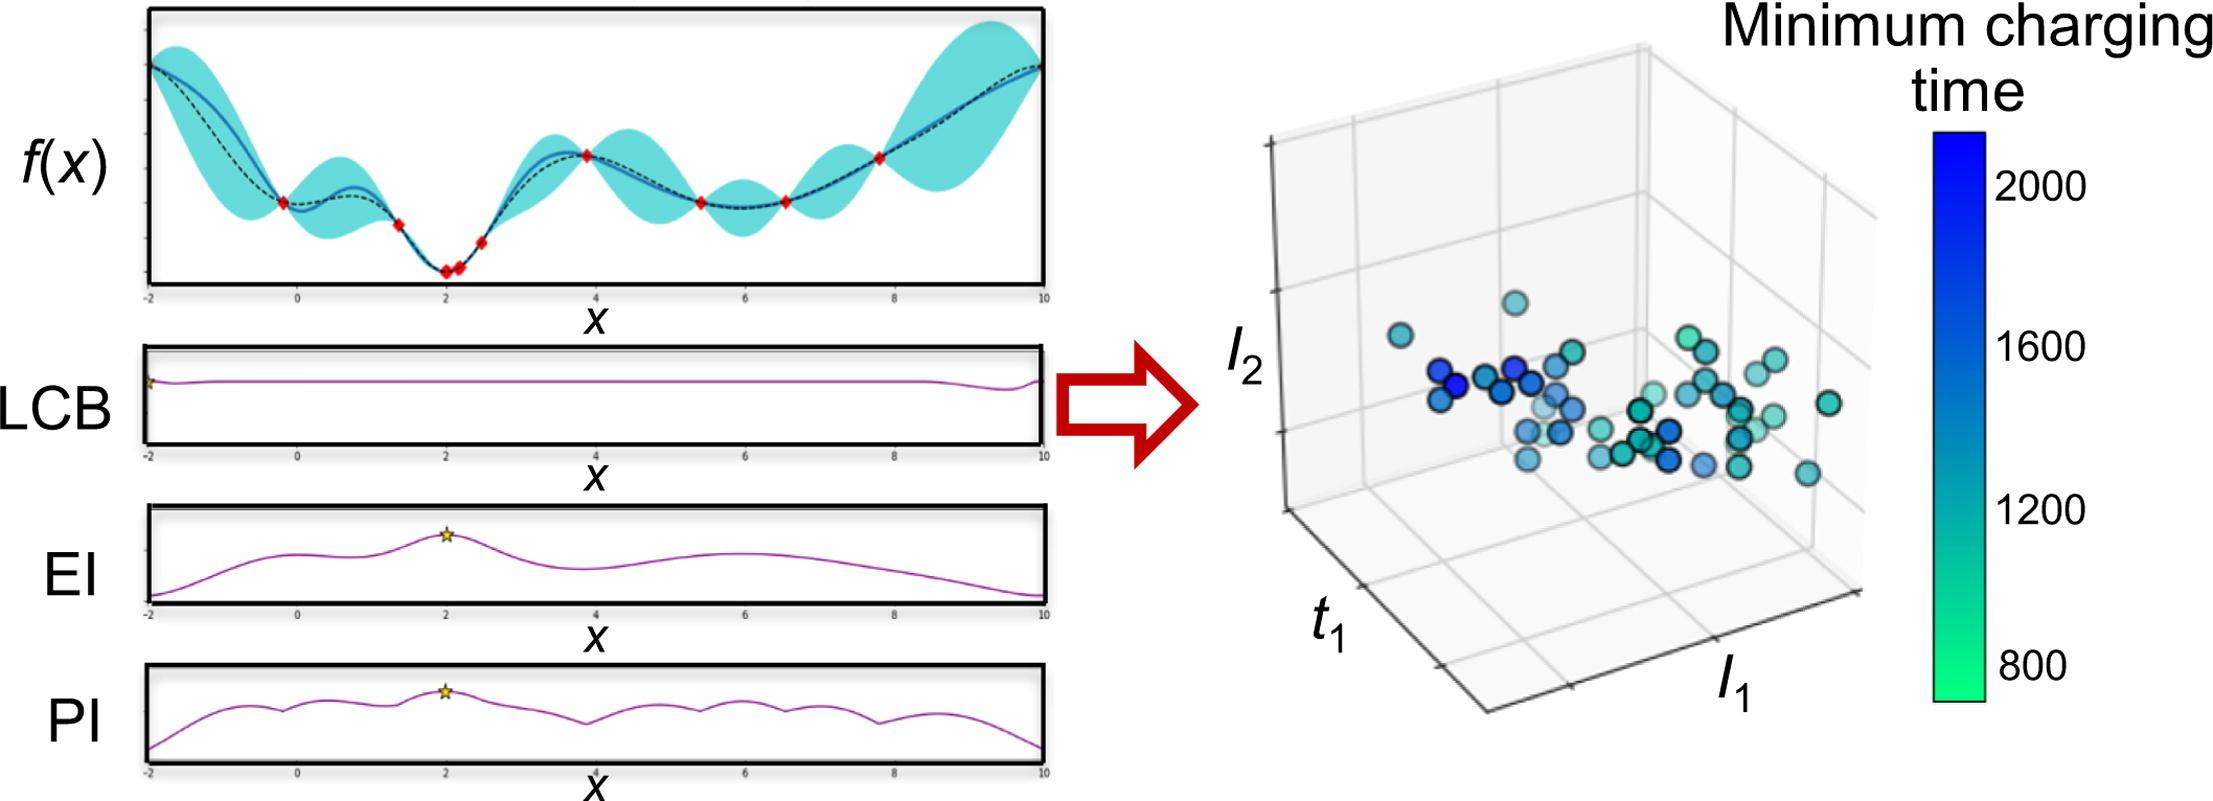
Lithium-ion batteries are one of the most commonly used energy storage device for electric vehicles. As battery chemistries continue to advance, an important question concerns how to efficiently determine charging protocols that best balance the desire for fast charging while limiting battery degradation mechanisms which shorten battery lifetime. Challenges in this optimization are the high dimensionality of the space of possible charging protocols, significant variability between batteries, and limited quantitative information on battery degradation mechanisms. Current approaches to addressing these challenges are model-based optimization and grid search. Optimization based on electrochemical models is limited by uncertainty in the underlying battery degradation mechanisms and grid search methods are expensive in terms of time, testing equipment, and cells. This article proposes a fast-charging Bayesian optimization strategy that explicitly includes constraints that limit degradation. The proposed BO-based charging approaches are sample-efficient and do not require first-principles models. Three different types of acquisition function (i.e., expected improvement, probability of improvement, and lower confidence bound) are evaluated. Their efficacies are compared for exploring and exploiting the parameter space of charging protocols for minimizing the charging time for lithium-ion batteries described by porous electrode theory. The probability-of-improvement acquisition function has lower mean and best minimum charging times than the lower-confidence-bound and expected-improvement acquisition functions. We quantify the decrease in the minimum charging time and increase in its uncertainty with increasing number of current steps used in charging protocols. Understanding ways to increase the convergence rate of Bayesian optimization, and how the convergence scales with the number of degrees of freedom in the optimization, serves as a baseline for extensions of the optimization to include battery design parameters. READ MORE
The modus operandi in materials research and development is combining existing data with an understanding of the underlying physics to create and test new hypotheses via experiments or simulations. This process is traditionally driven by subject expertise and the creativity of individual researchers, who “close the loop” by updating their hypotheses and models in light of new data or knowledge acquired from the community. Since the early 2000s, there has been notable progress in the automation of each step of the scientific process. With recent advances in using machine learning for hypothesis generation and artificial intelligence for decision-making, the opportunity to automate the entire closed-loop process has emerged as an exciting research frontier. The future of fully autonomous research systems for materials science no longer feels far-fetched. Autonomous systems are poised to make the search for new materials, properties, or parameters more efficient under budget and time constraints, and in effect accelerate materials innovation. This paper provides a brief overview of closed-loop research systems of today, and our related work at the Toyota Research Institute applied across different materials challenges and identifies both limitations and future opportunities. READ MORE
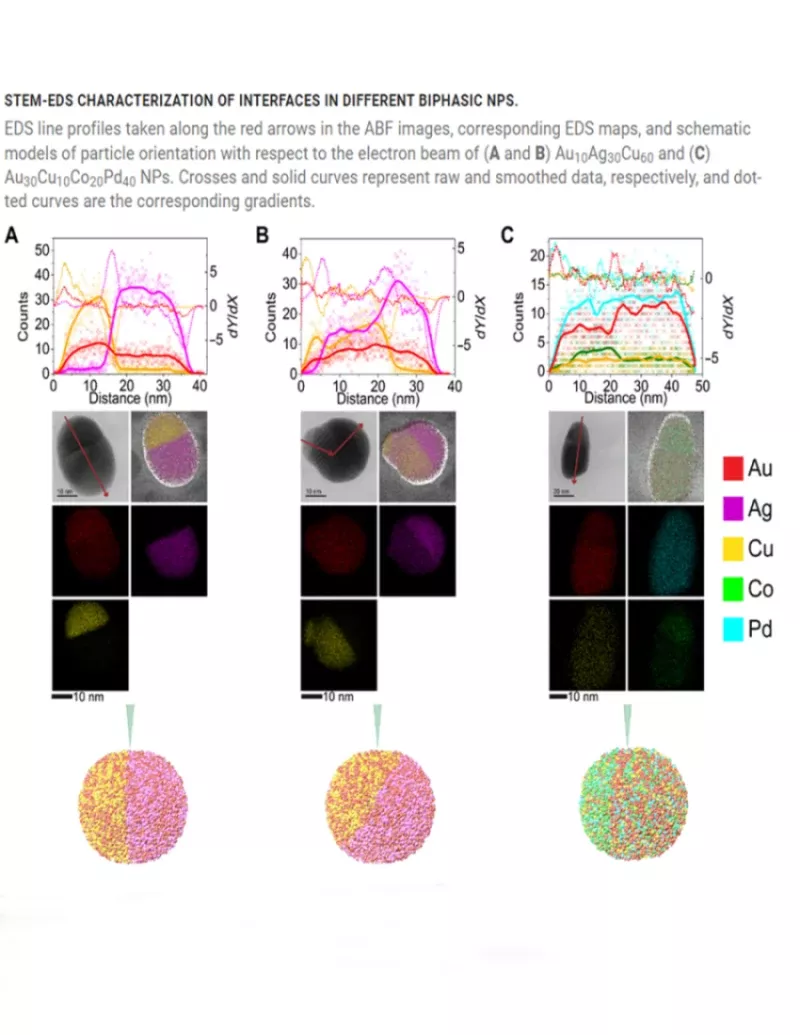
In materials discovery efforts, synthetic capabilities far outpace the ability to extract meaningful data from them. To bridge this gap, machine learning methods are necessary to reduce the search space for identifying desired materials. Here, we present a machine learning–driven, closed-loop experimental process to guide the synthesis of polyelemental nanomaterials with targeted structural properties. By leveraging data from an eight-dimensional chemical space (Au-Ag-Cu-Co-Ni-Pd-Sn-Pt) as inputs, a Bayesian optimization algorithm is used to suggest previously unidentified nanoparticle compositions that target specific interfacial motifs for synthesis, results of which are iteratively shared back with the algorithm. This feedback loop resulted in successful syntheses of 18 heterojunction nanomaterials that are too complex to discover by chemical intuition alone, including extremely chemically complex biphasic nanoparticles reported to date. Platforms like the one developed here are poised to transform materials discovery across a wide swath of applications and industries. READ MORE